Your file has vibrational data. At the moment, vibrations are handled by creating a short animation → out, back, in, back = 21 steps.
Before 2.0 is released, there is no great way to handle optimizations with a set of vibrations for each. Obviously, there are plenty of use cases for that, but it’s not currently supported. (I have a few ideas in the post-2.0, late-spring - summer - fall timescale.)
In the meantime, ORCA usually gives a trajectory XYZ file with all the energies as well.
The animation tool already appears to support keyboard control in version 1.103, provided that the toggle button is first activated (i.e., clicked to give it focus).
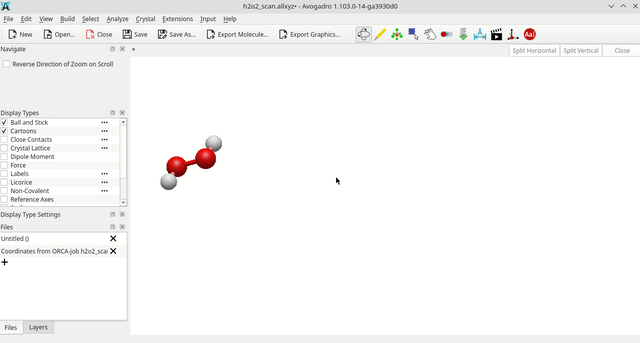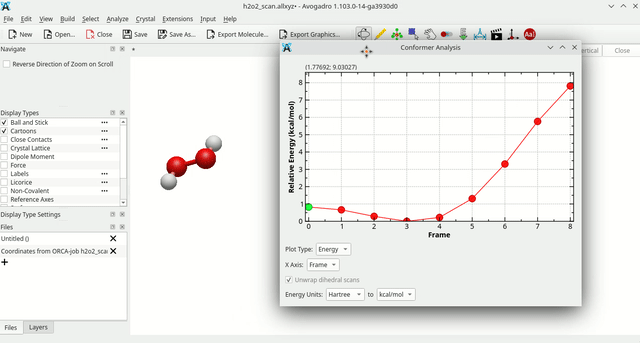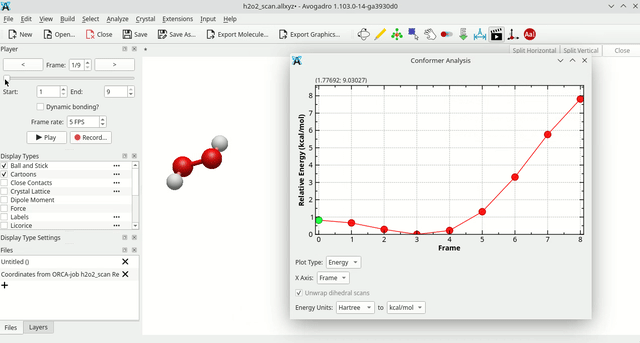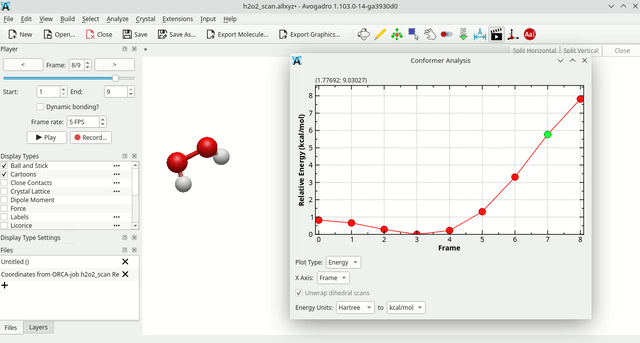
Moreover, I was actually working on this widget, adding point highlighting (among other things, aiming at plotting relaxed scans). My approach was similar to what indicated by @ghutchis, i.e. adding a new DataSeries that contains only the current point, with green color:
(in the animation, I first click on the toggle button to enable keyboard interaction and the points are explored with the keyboard)
I noticed that you had already implemented a highlighted point. Great!
However, the current approach seems to restrict the highlighted-point updates to interactions inside the conformer plot widget. In particular, when the active conformer is changed externally, for example through the animation tool (as shown in the GIF from the last post), the highlighted point is not updated with the code in the master branch (at least this is what I got). In my implementation, I managed to update it by calling updatePlot() from moleculeChanged() when the molecule emits an Atoms change. This works, but it also introduces extra lag because it recomputes properties and redraws the plot. While this is not noticeable for small conformer sets, it may become relevant for larger ones. Still, I think it is useful for the highlighted point to stay synchronized with the currently displayed conformer (maybe with a more efficient approach to replot the highlighted point only).
Yes. I’m just being extra-cautious with code changes right before the 2.0 release. There’s a real need for code to track the current conformer in the Molecule class itself.
Re-visiting this. Would it be possible to either add the keyboard shortcut directly to the Conformer Analysis window or else, updating the highlighted point as Javier was suggesting?
I will mention that now is a good time if you have particular properties (y-axis or x-axis) that you’d like to see in the conformer plot before v2.1 is released in October.
Hi. I would find it useful if there was an option (e.g. checkbox) to show absolute values, instead of relative ones. Namely, in my lecture, I am showing that both ground and first excited state of H2+ have an energy of -0.5hartrees at dissociation. It’d be nice to show it in the plot, while keeping the current default behaviour
Yes, sorry It is the time I am trying the program in my teaching and this came up. It is good anyway if this is added to the list of features for the next release. Thanks!

 It is the time I am trying the program in my teaching and this came up. It is good anyway if this is added to the list of features for the next release. Thanks!